all search
Tel: 0386-311-66695955, +86 18631137373
Factory Address: No.29-11 Jinyu Avenue, Zanhuang Economic Development Zone, Shijiazhuang City, Hebei Province, P.R.China
Office Address: 14th Floor, East Wing, Business Building 6, Xuhui Zhongrui Mansion, No.281 South Jianshe Avenue, Shijiazhuang City, Hebei Province
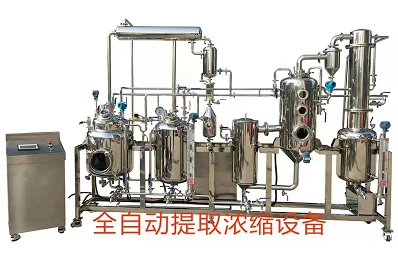
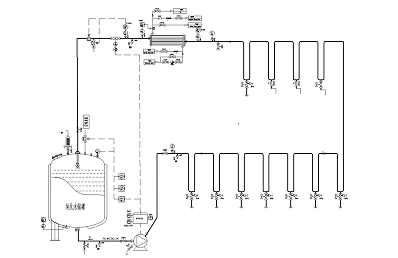
Fully automatic liquid dispensing systemIt is the core process equipment in the pharmaceutical production process, widely used in the concentration, dilution, and buffer preparation processes of injections, freeze-dried powder injections, oral liquids, biologics, eye drops, and other products. The material, structural design, self-control logic, and verification level of the equipment directly affect the quality and safety of drugs and also affect the pass rate of GMP on-site inspections.
Many pharmaceutical companies tend to focus only on production capacity and hardware pricing during the equipment procurement stage, neglecting key conditions such as regulatory compliance, process adaptation, and data integrity, resulting in a large amount of equipment modification, inability to complete verification, and inability to meet drug regulatory verification requirements after entering the site. This article combines the relevant requirements of drug production quality management standards, and summarizes the practical points of compliant selection of fully automatic liquid dispensing systems from the dimensions of process matching, hardware materials, sanitary structure, self-control system, verification documents, and supplier capabilities, providing reference for equipment procurement in pharmaceutical projects.
1、 Using our own production process as the basis for selection
Selection requires a clear understanding of product formulation, batch size, material characteristics, and cleanliness level requirements to determine the basic configuration of the liquid preparation system. Aseptic preparations and injectable products have higher requirements for airtightness, cleaning and sterilization capabilities; Ordinary oral preparations can adjust their configuration standards based on actual risk levels.
It is recommended to reserve reasonable redundancy for the effective capacity of the tank based on the production scale of a single batch, and it is not advisable to operate the tank at full capacity for a long time. High viscosity materials and suspension formulations need to be matched with corresponding mixing forms and speed ranges; Chemicals containing salt and organic solvents need to be evaluated for their chemical compatibility to avoid reactions between the materials and the components in contact with the equipment. Simultaneously confirm the cleanliness level of the equipment placement area and match the installation and operation conditions of the clean area.
2、 Compliance verification of material and surface treatment of contact material components
The tanks, pipelines, valves, and mixing components that come into direct contact with the medicinal solution are the fundamental components of compliance. The mainstream choice for pharmaceutical liquid preparation systems is 316L sanitary grade stainless steel. The supplier is required to provide material certification and spectral testing reports to confirm the authenticity of the material.
The polishing treatment of the inner surface of the equipment needs to meet the standards of the pharmaceutical industry, with smooth transitions of the inner wall welds, no pits, burrs, or dead corners, to reduce the risk of material hanging on the wall. Priority should be given to using sanitary grade connection methods for pipelines to reduce dead corners and blind pipes. Non metallic components such as seals and diaphragm valve gaskets need to have corresponding material compliance certificates to meet the requirements for drug contact and use, and to avoid quality risks caused by precipitates.
3、 Evaluation of Health Structure and CIP/SIP Functional Configuration
The fully automatic liquid preparation system requires hardware conditions that can achieve online cleaning CIP and online sterilization SIP, and is an important configuration for sterile pharmaceutical production lines. The layout of the tank spray ball needs to cover all areas inside the tank to ensure no blind spots during cleaning; The layout of pipeline valves reduces hygiene blind spots, making it easier to complete cleaning and sterilization verification.
The system needs to be equipped with sterile respirators and compliant sampling valves to meet the requirements of production sampling and pressure balance inside the tank. Priority should be given to selecting structures that reduce the risk of leakage and minimize contamination hazards at the shaft seal position. When switching production between different varieties, relying on a well-established CIP process to reduce the risk of cross contamination, the selection stage needs to evaluate whether the cleaning and sterilization plan is suitable for actual production conditions based on the frequency of product switching.
4、 Automation control system and data integrity requirements
The automatic control software of the fully automatic liquid dispensing system needs to meet the relevant requirements of GMP on data integrity, which is also a focus of drug regulatory verification. The system should have hierarchical permission management, with corresponding operational permissions assigned to different positions to achieve controllable operational behavior. Supporting electronic signature and audit tracking functions, key operations, parameter modifications, and alarm information can be fully recorded and retained, and data cannot be tampered with at will.
The system needs to implement formula management function, which can store multiple sets of production formulas and automatically complete processes such as feeding, mixing, temperature control, and conveying; temperature pH、 Key process parameters such as conductivity, liquid level, and weighing can be collected in real-time, automatically saved, and generate traceable batch reports. Simultaneously evaluate the compatibility between the equipment and upper level systems such as MES to meet the requirements of digital production construction in the factory. The supporting instruments need to support calibration to ensure accurate and reliable detection data.
5、 Is the supporting verification file complete and implementable
Equipment hardware compliance does not necessarily mean project compliance. A complete verification document is a necessary condition for the equipment to be put into formal production. During the selection phase, it is necessary to confirm that the supplier can provide technical document templates related to DQ design confirmation, IQ installation confirmation, and OQ operation confirmation to support the enterprise in completing PQ performance confirmation work in the later stage.
The relevant documents include equipment design specifications, material certificates, welding inspection records, polishing inspection reports, instrument calibration certificates, software instructions, alarm lists, and other materials. The more complete the documentation, the more advantageous it is for enterprises to carry out verification work, reduce the workload of later data supplementation, and better cope with GMP inspections.
6、 Key points of supplier comprehensive capability evaluation
In addition to the hardware parameters of the equipment, the supplier's experience in pharmaceutical industry projects is equally important. Prioritize investigating the implementation cases of suppliers in similar dosage form projects, understand their understanding of pharmaceutical processes and GMP regulations, and avoid manufacturers who only have experience in manufacturing general chemical equipment.
Simultaneously evaluate the supplier's design, processing, and on-site debugging capabilities, and confirm that they can complete on-site installation and debugging, personnel training, and post technical support. Understand the spare parts supply cycle to facilitate equipment maintenance in the later stage. Don't just rely on equipment quotations as the basis for selection decisions, comprehensively weigh hardware configuration, compliance support, and overall level of technical services to reduce the risk of project renovation in the later stage.
7、 Common risk avoidance during the selection stage
1. Don't just look at the appearance of the equipment, check material reports, polishing tests, welding records and other supporting documents to prevent low standard materials from being used in pharmaceutical production. 2. Do not underestimate software compliance. Functions such as audit tracking, permission management, and electronic signatures need to be clearly written into the requirements during the procurement phase, and the cost of later transformation is relatively high. 3. Fully consider product switching and multi batch production scenarios, evaluate the actual operational effects of CIP and SIP, and avoid situations where cleaning validation is difficult to pass in the later stage. Fourthly, confirm the delivery scope of verification documents in advance to avoid missing key technical information after arrival.
Summary
pharmaceutical industryFully automatic liquid dispensing systemCompliance selection requires comprehensive evaluation based on six dimensions: process condition matching, material and surface treatment, sanitary grade structure and CIP/SIP configuration, self-control and data integrity, complete set of verification documents, and supplier industry experience. Equipment procurement cannot simply refer to general chemical equipment standards, comply with GMP regulations, avoid material contamination, cross contamination, data non traceability and other risks from the source, ensure stable and controllable drug production processes, and meet the dual requirements of production operation and drug supervision verification.